
 Repository
Current collected state Current
Observed since 20 Oct, 17:21 UTC
No superseded metadata states collected. Unchanged observations do not create new revisions.
ABFE Workflow is a pipeline variant of our prior work.
WARNING: The repo is currently under development - No Guarantees at the moment. But will be soon at a first version. This goes with a manuscript, that is currently submitted. :)
A snakemake based workflow for ABFE calculations using GMX. The workflow can be scaled on Slurm Queuing systems. The here provided Cyclophilin D Test systems and experimental values originate from:
Here a visualization of the triggered process:
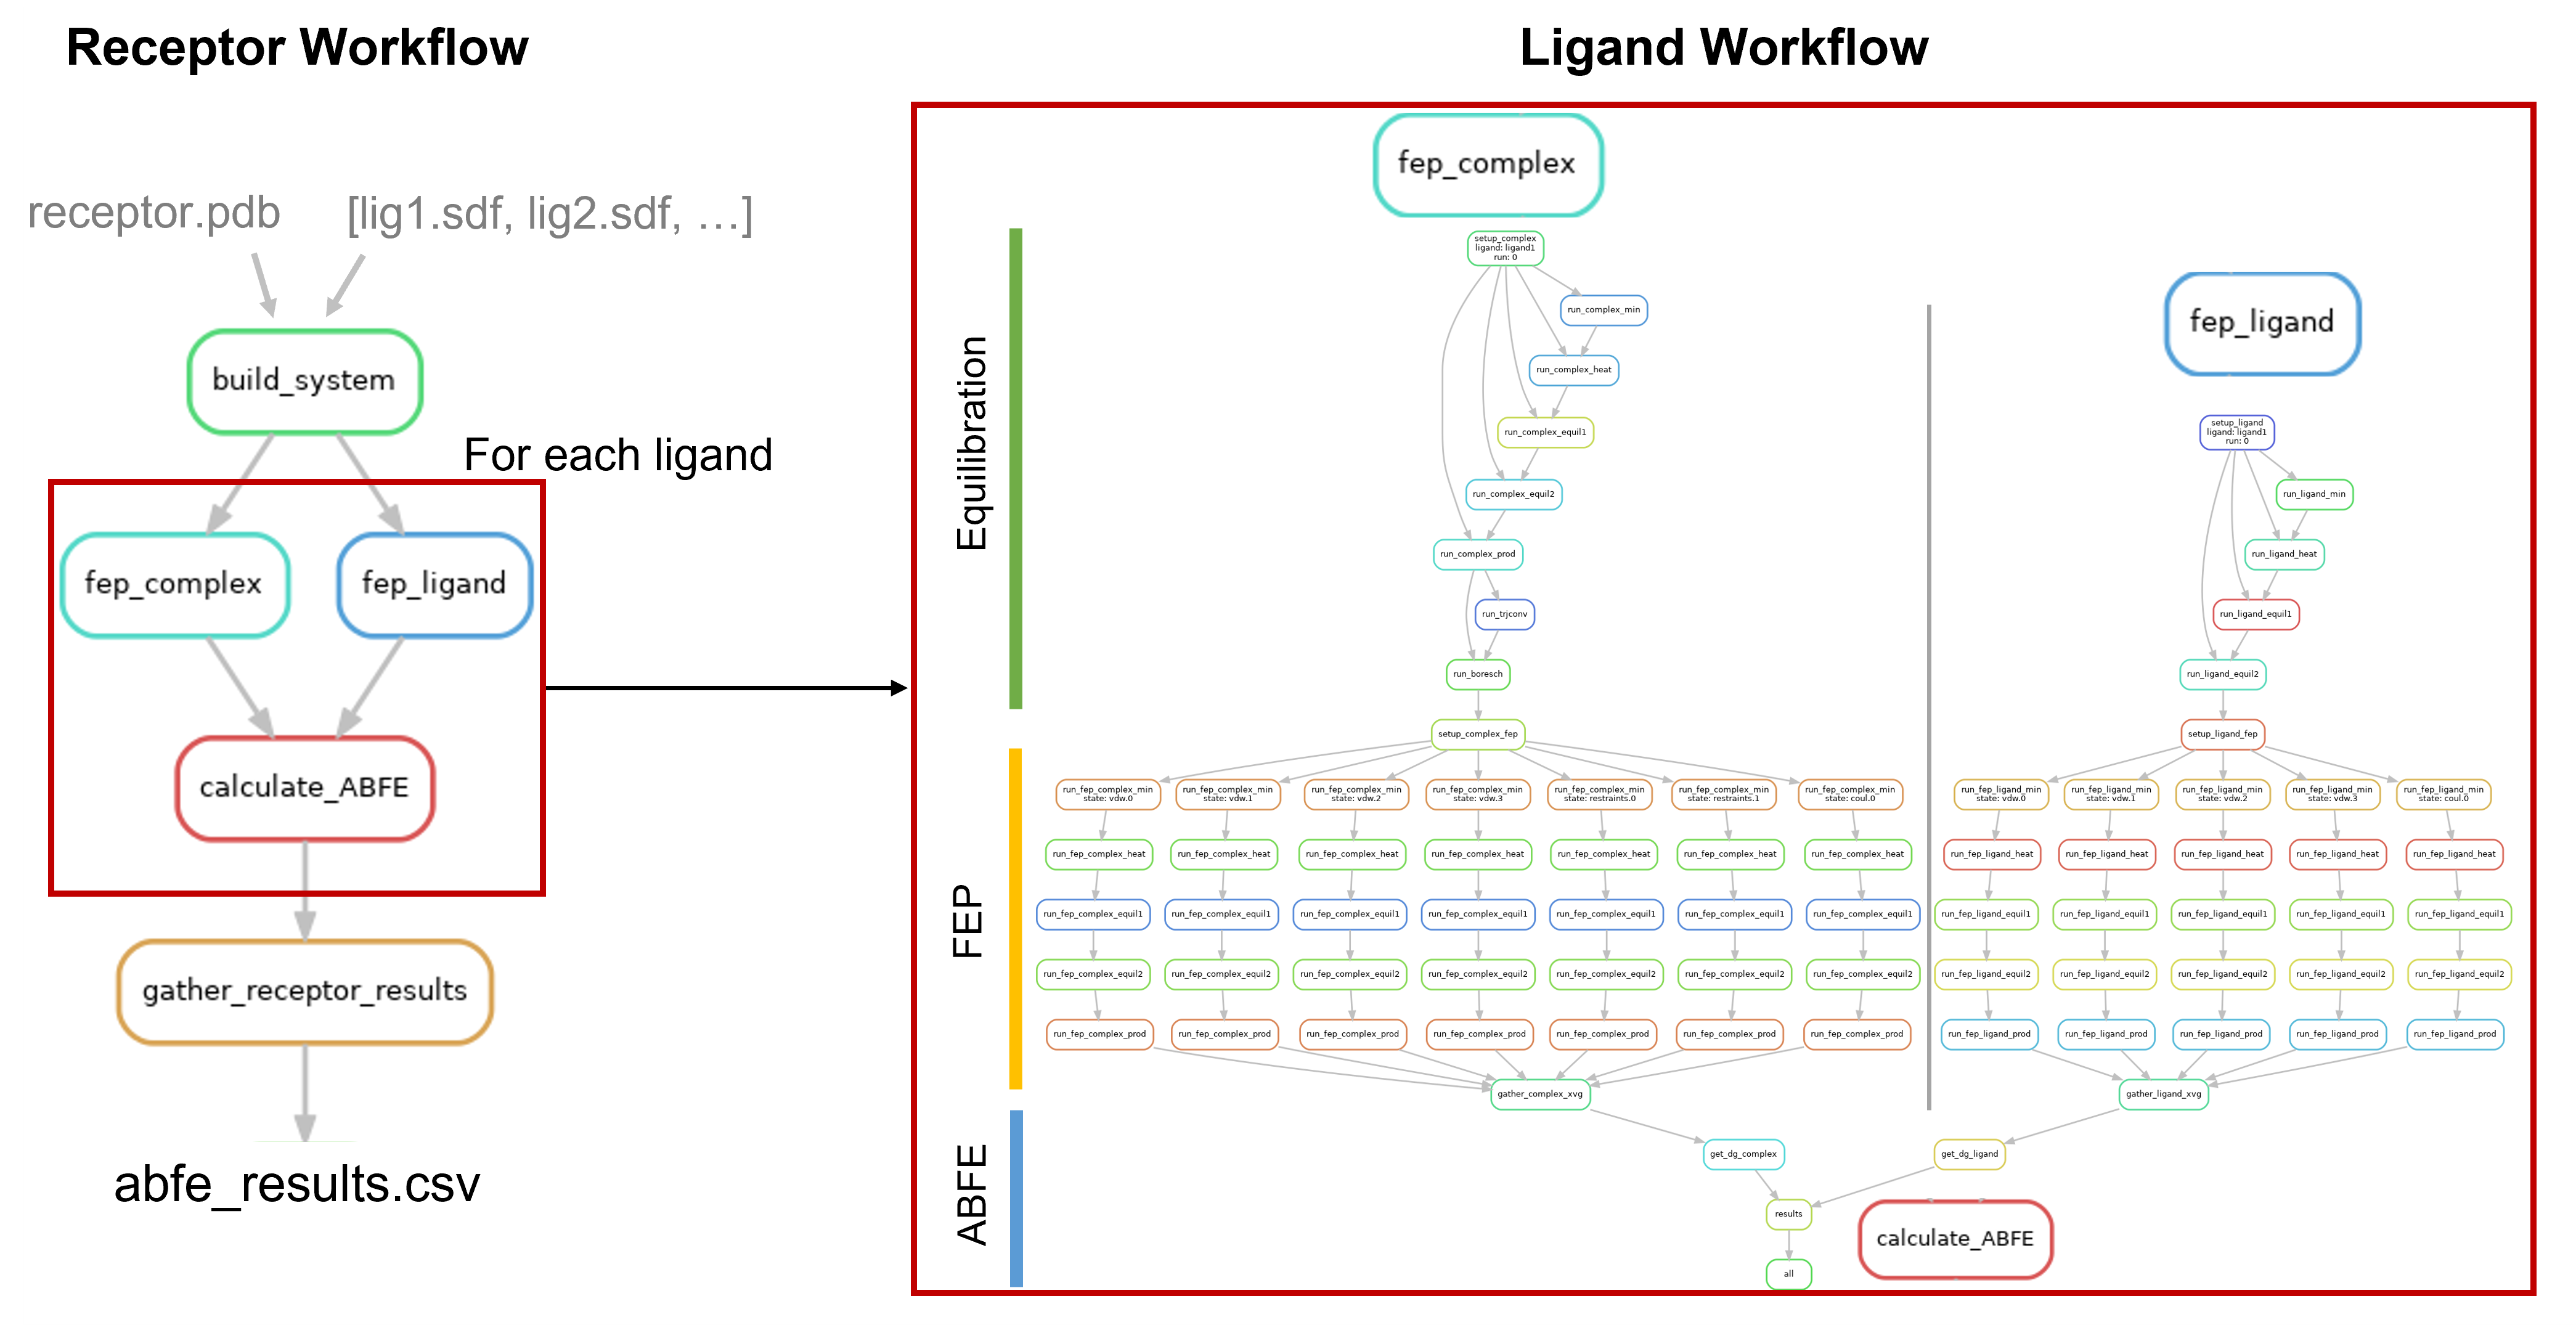
We are currently improving the user experience. The input was simplified to simply only the need of providing a pdb file for the receptor and .sdf files for the ligand.
>cli-abfe -h

usage: cli-abfe [-h] -p PROTEIN_PDB_PATH -l LIGAND_SDF_DIR -o OUTPUT_DIR_PATH [-c COFACTOR_SDF_PATH] [-nc NUMBER_OF_CPUS_PER_JOB] [-nj NUMBER_OF_PARALLEL_JOBS] [-nr NUMBER_OF_REPLICATES] [-submit]
                       [-gpu] [-hybrid]

optional arguments:
  -h, --help            show this help message and exit
  -p PROTEIN_PDB_PATH, --protein_pdb_path PROTEIN_PDB_PATH
                        Input protein pdb file path
  -l LIGAND_SDF_DIR, --ligand_sdf_dir LIGAND_SDF_DIR
                        Input ligand(s) sdf file path
  -o OUTPUT_DIR_PATH, --output_dir_path OUTPUT_DIR_PATH
                        Output approach folder
  -c COFACTOR_SDF_PATH, --cofactor_sdf_path COFACTOR_SDF_PATH
                        Input cofactor(s) sdf file path
  -nc NUMBER_OF_CPUS_PER_JOB, --number_of_cpus_per_job NUMBER_OF_CPUS_PER_JOB
                        Number of cpus per job
  -nj NUMBER_OF_PARALLEL_JOBS, --number_of_parallel_jobs NUMBER_OF_PARALLEL_JOBS
                        Number of jobs in parallel
  -nr NUMBER_OF_REPLICATES, --number_of_replicates NUMBER_OF_REPLICATES
                        Number of replicates
  -nosubmit             Will not automatically submit the ABFE calculations
  -nogpu                don't use gpus for the submissions?
  -nohybrid             don't do hybrid execution (complex jobs on gpu and ligand jobs on cpu (requires gpu flag))
An example usage is provided with the examples/example_execution.sh, that uses the ABFE_Calculator.py script.
If you remove the submit flag, you can a start a run, that only sets up the folder structure. (checkout our example folder)
Additional script information is provided via:
  conda activate abfe

  cli-abfe -h
  # or
  cli-abfe-gmx -h
Running an ABFE Campaign from Bash:
  conda activate abfe
  cli-abfe -p <path>/receptor.pdb \
           -l <path>/myligands \
           -o <path>/Out  \
           -nogpu -nohybrid -nc 8
The input is suggested to be structured as follows for the commandline option:
For the python call:
Alternativley you can provide gromacs input files and use the command line tool ABFE_GMX_CLI. Please make sure your ligand is called LIG in the gmx files. The input sturucture should look like this:
The package can be installed like the following script:
  cd ABFE_workflow
  conda env create --file ./environment.yml
  conda activate abfe
  pip install .
Download counts and popularity will appear here as the collector receives them.
| Version | Published | Direct dependencies | Open version |
|---|---|---|---|
| 0.5.0 | 5 Oct 2023 | 9 |
This describes our collected data; it is not a security assessment.
Observed since 20 Oct, 17:21 UTC
No superseded metadata states collected. Unchanged observations do not create new revisions.
package:10ee57b8b5ae1ead19e95d89e03db1ff0728d918b611a15cf9d4eee80ceaee64